
 1
 0
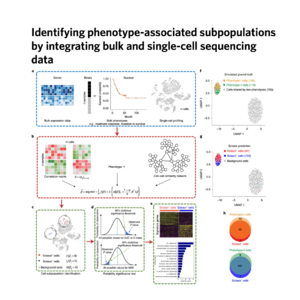
The paper introduces Scissor, a novel computational tool designed to identify specific cell subpopulations by integrating single-cell RNA sequencing with bulk tissue data and clinical phenotypes. While single-cell technologies excel at characterizing tissue complexity, they often lack the statistical power to link cell clusters to specific clinical outcomes like survival rates, drug responses, or disease stages. Scissor overcomes this limitation by quantifying similarities between individual cells and large-scale bulk datasets, such as those from The Cancer Genome Atlas (TCGA), to pinpoint the most disease-relevant subsets. The researchers validated the tool across diverse conditions, successfully identifying aggressive lung cancer cells linked to hypoxia, melanoma T cells associated with immunotherapy success, and specific cell types involved in Alzheimer’s disease. Ultimately, this framework provides a flexible, hypothesis-free approach for discovering prognostic biomarkers and potential targets for cell-directed therapies.
References:
- Sun, D., Guan, X., Moran, A.E. et al. Identifying phenotype-associated subpopulations by integrating bulk and single-cell sequencing data. Nat Biotechnol 40, 527–538 (2022). doi.org
